UFF and UFF4MOF
xtalforge-ff implements the Universal Force Field natively. The parameter table is data/uff.json: 218 rows ported from Crystal-Builder xtal/ff/uff/params.py (Rappé 1992, Addicoat 2014, Coupry 2016), with the reference units (Angstrom, degrees, kcal/mol). The public boundary converts to eV and eV/Angstrom.
Typing
An atom type is (element, geometry, oxidation state). The typer combines the element table, the perceived bond graph and the local geometry:
- explicit rules for H, C, N, O, F, Cl, Br, I, S, P and the common metals;
- aromatic ring bonds use the resonant carbon/nitrogen types;
- terminal atoms refine their geometry from the number of neighbours;
- framework metals prefer the UFF4MOF rows when the set is
Uff4Mof; - a dummy
Xatom has no parameters and never contributes.
Typing records the chosen AtomType and a Confidence, so a caller can see why an atom got a type. UffModel exposes the prepared topology.
Energy terms
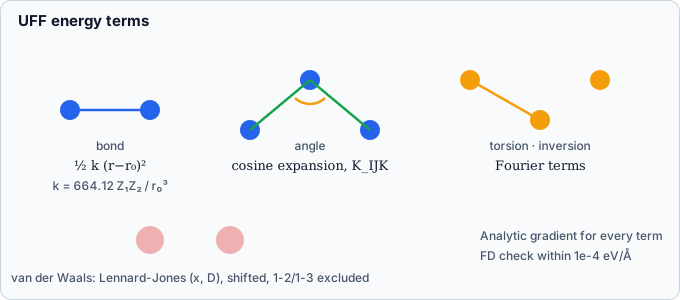
- Bond:
0.5 k (r - r0)^2,k = 664.12 z1 z2 / r0^3. - Angle: the reference cosine expansion whose curvature at
theta0is the UFFK_IJK. - Torsion and inversion: the reference Fourier terms.
- van der Waals: Lennard-Jones with the UFF
xandDparameters, 1-2 and 1-3 exclusions, a cutoff and a shifted potential.
Every term has an analytic gradient. The finite-difference test moves each coordinate by a small Cartesian step and compares forces within 1e-4 eV/A.
Validation
- Ethylene energy
0.1113230794 eV, matching the reference term-by-term sum. - Diamond periodic energy
1.7200457169 eV. - Argon dimer shifted well
-0.0080043731 eV. - Methane at its UFF bond length, water at its angle, a benzene resonant ring and a Zn4O UFF4MOF node.
The Crystal-Builder golden comparison over the full corpus is not run in CI; the values above are the regression anchors.